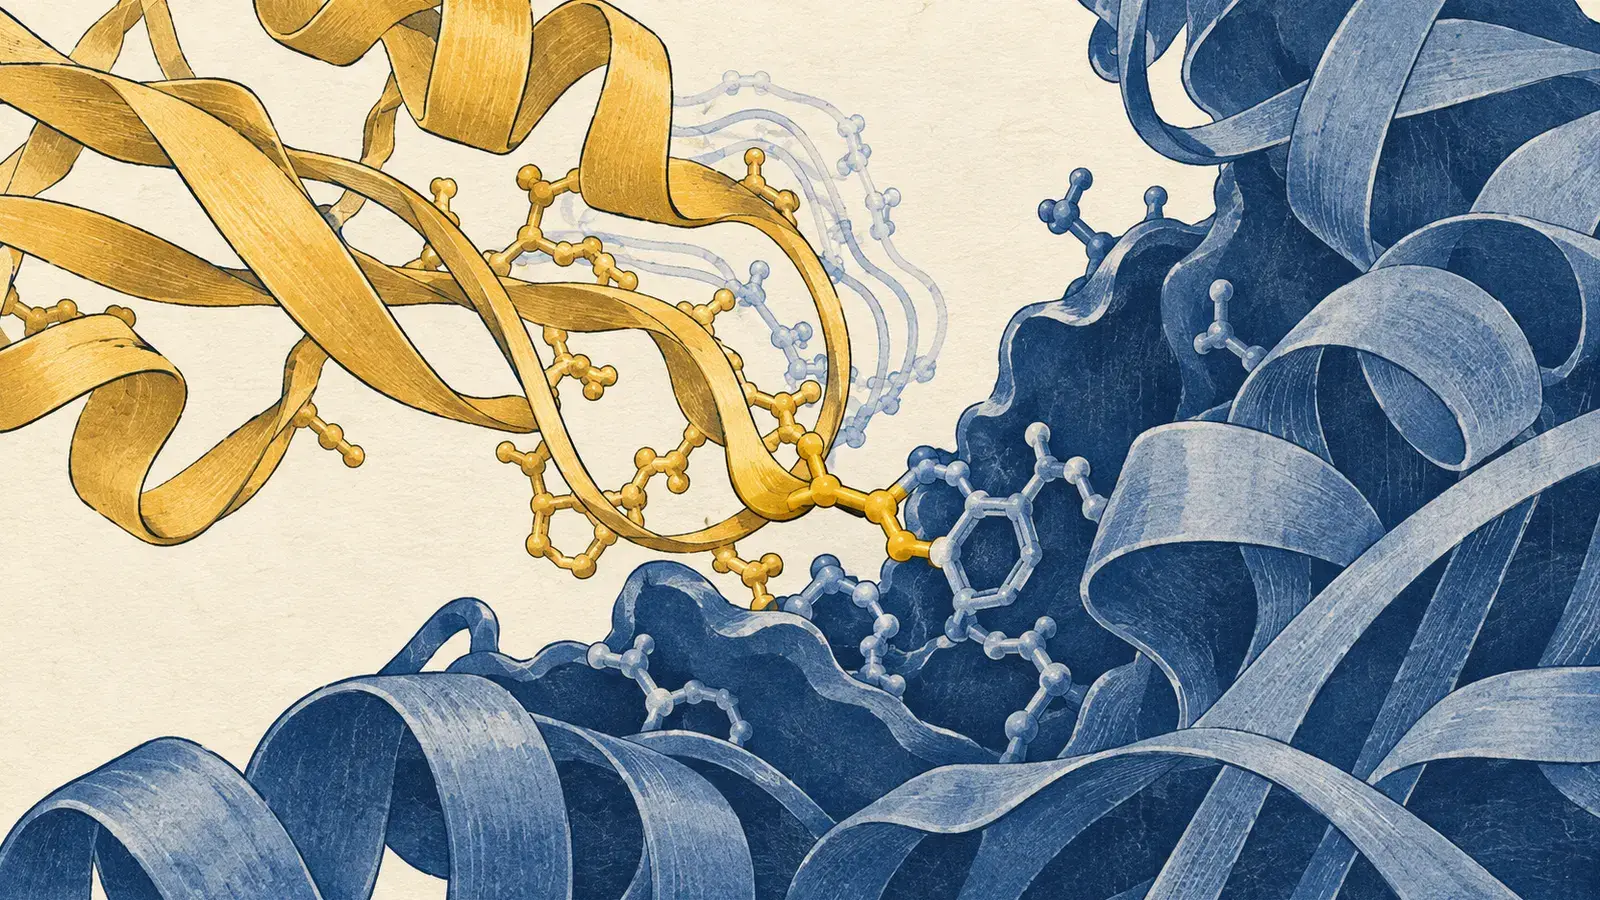
A new computational platform, DyME, automates the study of how changes to amino acids affect recognition and binding between biomolecules. The paper, published on October 7, 2026, in PLoS Computational Biology, presents a single environment for creating protein variants, simulating them and comparing the results. The platform is designed to organize up to thousands of molecular systems. According to the researchers, the aim is to reduce the manual work that makes large-scale studies difficult.
Molecular recognition depends on the characteristics of the regions where two molecules come into contact. By replacing selected amino acids, researchers can explore changes in binding stability or in a protein’s preference for different molecular partners. Methods that examine static structures struggle to capture the full complexity of these interactions. Molecular dynamics simulations track the behavior of systems at the atomic level, but each distinct variant needs its own simulation. Preparing and comparing hundreds of variants therefore creates a substantial practical workload.
DyME starts with the three-dimensional structure of a complex containing a protein and another protein, a peptide or DNA. The user specifies which molecule will be modified, and the platform suggests amino acid positions at the interface for further investigation. These suggestions can be changed, and other positions of interest can also be selected. The desired substitutions and groups of positions to be examined together are then specified. These choices are used to create a library of variants with single, double and triple changes.
For each variant, the system automatically builds the three-dimensional structure and prepares the files needed for the simulation. It uses Modeller for structural changes, Amber file conventions and OpenMM for molecular dynamics simulations in a solvent environment. Jobs are distributed across available graphics processing units, with each unit running one simulation at a time. The distributed architecture allows computing nodes to be added across more servers. Systems with nonstandard components, such as synthetic amino acids, are also supported through appropriate parameters.
After each simulation is complete, automated processes extract information about structural behavior, binding free energies, contacts between molecules and water around the interaction regions. The processed data are collected in a central MongoDB database, while the raw files are kept separately. This organization allows searches combining multiple criteria and comparisons of features across many simulations, without having to manage each result individually. The database also tracks job status and coordinates successive processing stages. Users can explore results that are already available before the entire library is complete.
Access is through a web interface, with tools for numerical comparisons, graphs and three-dimensional viewing. Variants can be ranked by their calculated binding energies and compared with the original reference structure. Other graphs show structural deviations during the simulation and the energy contribution of selected amino acid positions. The interface allows multiple variants to be viewed at once, as well as graphs and numerical data to be exported. Users can also add a combination of changes missing from the library or request additional simulation repeats.
Two specialized tools focus on aspects of molecular recognition. Water-site Explorer helps study water sites at the interface, as water molecules can participate in interactions between biomolecules. Specificity Finder compares two projects in which the same molecule being modified binds to different molecular partners. Its purpose is to identify changes that strengthen binding to one partner and weaken it to the other, supporting the investigation of selectivity.
To evaluate the platform, the researchers presented a case study based on available experimental data. They report that the binding free energies calculated through DyME correlated with experimental dissociation constants, which describe molecular binding. Water-site mapping also highlighted water-mediated interactions that had been identified experimentally. The authors present these findings as evidence of the tool’s usefulness for the rational design of protein variants.
DyME’s source code is openly available on GitHub, while the study’s simulation trajectories and associated input files are available on Zenodo. The platform runs on Linux and is distributed through Docker and Apptainer containers to make its individual components easier to install. The study is open access, and the authors state that they received no specific funding for the work and have no conflicts of interest.
Comments